Why Is Sec Kennedy & FDA Still Hiding The DANGEROUS UNDISCLOSED DNA In Every Covid Vial?
Part 5 of 7 — Independent labs confirmed it. Health Canada acknowledged it. Shall we all finally force the answer and removal from market? Federal mandamus files next week to make Kennedy do DUTY!
Interest of Justice — April 17, 2026 — Citizen Petition Series, Part 5 of 7 — Docket FDA-2025-P-1807 (click to open the public docket)
THIS IS PART 5 OF 7. The full series walks through Citizen Petition FDA-2025-P-1807 section by section. FDA has been silent for 305 days — 125 days past the statutory response deadline.
IF YOU’RE NEW HERE
IoJ filed a citizen petition — FDA-2025-P-1807 — and the FDA is lagging on it. Hard. They had 180 days to respond. The deadline was December 13, 2025. We are now 305 days in with no substantive answer.
We are about to file a federal mandamus in the U.S. District Court for D.C. to force FDA to answer — and along the way, to demand the Congressional report the Department of Defense was legally required to produce before any bio-agent testing on civilians. That report does not exist. FOIA confirms it.
IoJ is reader-supported. No corporate sponsors, no law firm. The petition runs well over 130 pages of primary filing plus hundreds more pages of exhibits — because FDA itself told petitioners to pack everything into one document. The petition asks the agency to do what it should have done years ago — reclassify mRNA and adenoviral vector products as gene therapy, impose a clinical hold, and fix the record.
This is a seven-part series walking through the petition section by section. Whichever part you landed on, the others are linked below.
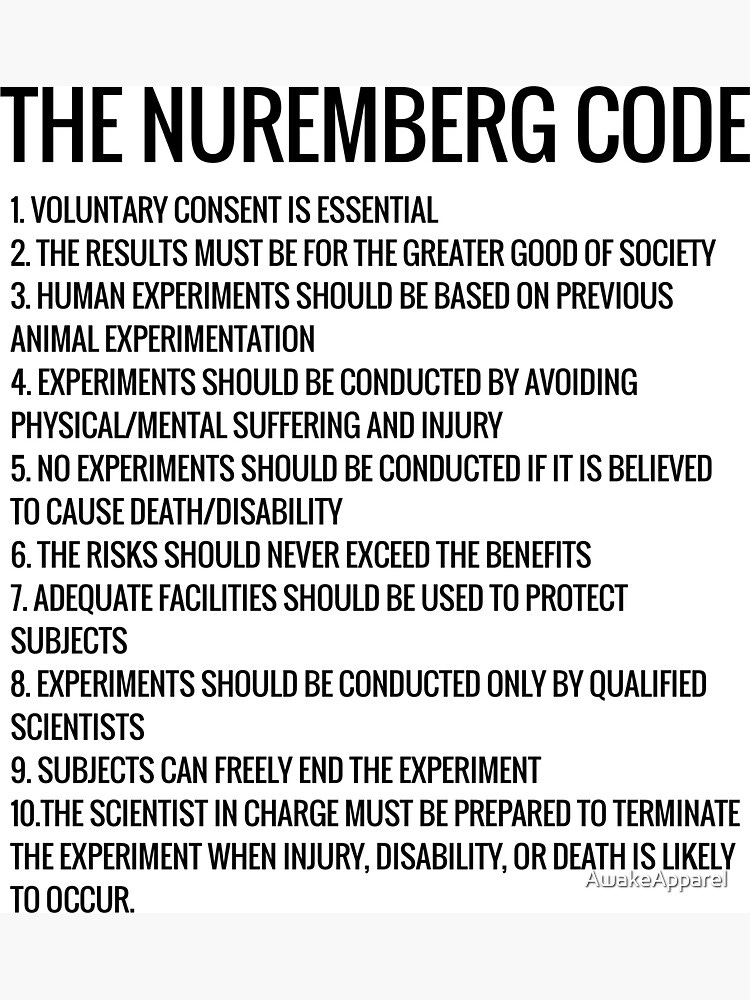
We know most people literally don’t care and are burnt out, BUT its important… so we are putting out this series to educate everyone on the details & preparing for the legal battle that will mark humanities turn back to Nuremberg Code & ethics.
Part 5 — The Adulteration and SV40 Problem (you are here)
Part 6 — The International Law Dimension (coming soon)
Part 7 — The Remedy We Are Asking For (coming soon)
TIRED OF WATCHING FDA LET PFIZER SLIDE?
You are not the only one. For years the findings have been in the peer-reviewed record and sister regulators have confirmed them — and FDA has said nothing. That is what ends with our court case being filed next week.
Help us stop the shots! Share, comment and donate now!
WHY HAS NO FEDERAL AGENCY ANSWERED FOR THE DNA THAT IS SO DANGEROUS?
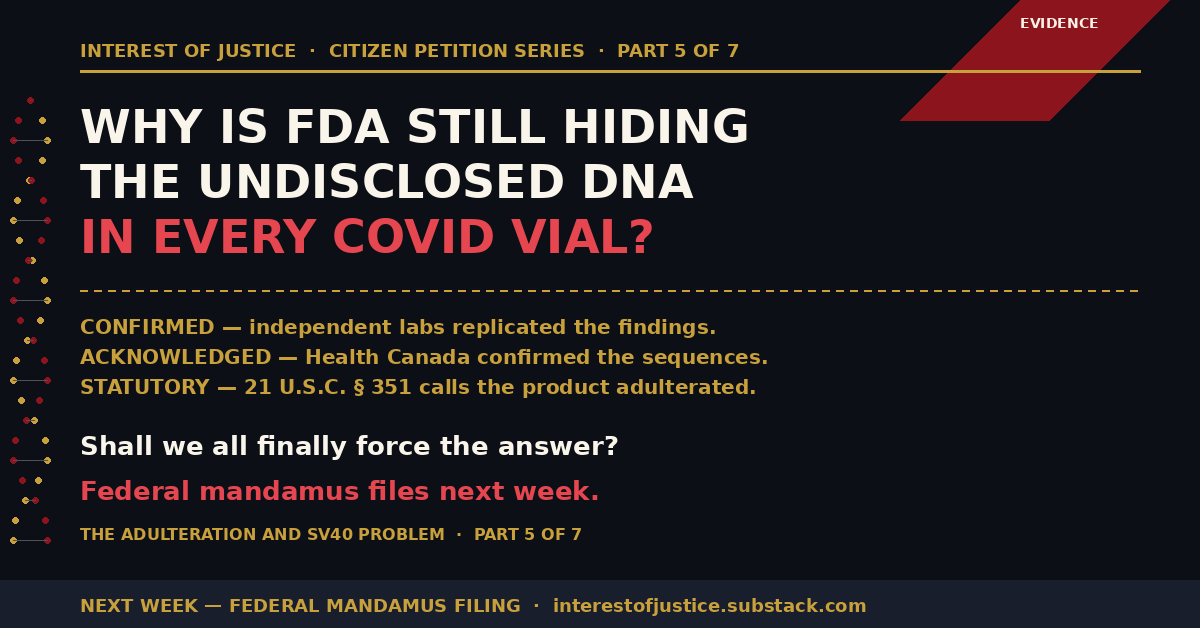
The evidence is already on the record. Three things have been established outside FDA —
CONFIRMED — Independent laboratories replicated Dr. Janci Lindsay’s SV40 findings. Kevin McKernan, formerly of the Human Genome Project sequencing team, did independent sequencing with the same result.
ACKNOWLEDGED — Health Canada publicly confirmed the undisclosed DNA sequences in Pfizer vials. That is a sister-regulator admission on the record.
STATUTORY — 21 U.S.C. § 351 calls a product adulterated when its actual composition differs from its label. The statute is already met.
TIMESTAMPED FOR YOU TO SEE HIM CALL SV40 “one of the most carcinogenic materials that is known to man”
IN THE VIDEO BELOW SEC KENNEDY AGREES - SO WHY IS HE LAGGING?
DIRECT QUOTE IN VIDEO ABOVE: “The polio vaccine contained a virus called simian virus 40, SV40. It’s one of the most carcinogenic materials that is known to man. In fact, it’s used now by scientists around the world to induce tumors in rats and guinea pigs in labs.”
He added that 98 million Americans received the contaminated polio vaccine. Two years after that statement, independent laboratories confirm SV40 promoter sequences in every COVID mRNA vial tested. Dr. Janci Lindsay testified under oath to the South Carolina Senate about exactly that — same SV40 class of contamination, now in a product deployed to 670 million Americans.
Kennedy — the Secretary of HHS who said SV40 is “one of the most carcinogenic materials known to man” — has done nothing. Let’s make him act.
No public statement. No written directive to FDA. No clinical hold. No recall. No correction of the biologics record. The petition documenting SV40 in every vial has been on his desk for 305 days. This is NOT the only petition on his desk about SV40.
He knows what is in the product. He said so himself. And he has not used the authority or performed the duty his own office requires.
So next week we have no choice but to file — to force it through.
This is the part that stings. Robert F. Kennedy Jr. — now Secretary of Health and Human Services — spent decades publicly questioning vaccine safety. He is named in the mandamus caption. He holds plenary authority and a non-discretionary duty under 42 U.S.C. § 262 to correct the biologics record and direct FDA’s response. The statute uses the word “shall,” which the D.C. Circuit held in Cutler v. Hayes creates “a consequential obligation on the agency, barring textual or contextual indications to the contrary.” That is not discretion. That is a duty the Secretary is legally required to perform. The petition has been on his desk for 305 days. The findings are already in the peer-reviewed record. The FOIA evidence on the DoD predicate is already in hand. He has everything he needs.
And still — no action. No public statement. No written direction to FDA. The Secretary who built his reputation on exactly this kind of accountability has not used the authority or fulfilled the duty his own office gives him.
We MUST file — so the court can make him finally take action.
The Secretary of HHS has the authority and the duty to act. He has held both for 305 days and chosen silence. Mandamus lies to compel a duty — not discretion. Next week we file so the court can make him use the authority and perform the duty he has been refusing to.
IF FDA DOES NOT DISPUTE THE FINDINGS — WHY ISN’T THIS URGENT? Sigh…
This is the part that should break the silence on its own. FDA has had years to put a counter-finding on the record. If the agency had data refuting Dr. Lindsay’s SV40 sequences, or Kevin McKernan’s independent sequencing, or Health Canada’s public acknowledgment, FDA would have said so — loudly, with methodology, with counter-sequencing results. That is how regulators defend product composition when they actually disagree.
FDA has not disputed any of it. No counter-finding. No written disagreement. No explanation. Silence.
An undisputed finding means the statutory definition of adulteration is already met. And an adulterated biologic deployed to 670 million Americans should be, under the agency’s own rulebook, a Grade A emergency — clinical hold, recall, investigation, accountability. It has not been treated as one. Not by FDA. Not by HHS. Not by the Secretary who has the authority to order it.
Either FDA disputes the findings — in which case the agency must put that dispute on the public record. Or FDA accepts them — in which case the agency must act. Silence is not an option the statute permits.
We have added multiple issues in the Petition that once answered will absolutely force the the Agency to stop the shots once and for all.
YOU CAN HELP FINALLY HOLD FDA ACCOUNTABLE. HELP US STOP THE SHOTS!
Here is what this part is about, in plain English. Federal law at 21 U.S.C. § 351 calls a drug adulterated when its actual composition differs from what the label says. Every COVID mRNA vial independent laboratories have tested contains residual DNA fragments that include SV40 promoter sequences. None of that was on the label. None of it was in the informed consent material. Health Canada has acknowledged the findings. Kevin McKernan’s independent sequencing replicated them and Dr. Janci Lindsay’s toxicology team documented them. Under § 351 — and under 18 U.S.C. § 178’s definition of biological agent as something capable of causing death, disease, or biological malfunction — the product meets multiple federal statutory definitions that should have triggered enforcement action years ago. Next week a federal mandamus asks the D.C. District Court to order FDA to answer for that.
The long version — with the sequencing evidence and the statutory citations — is below.
FOR THE DIEHARDS — THE FULL LEGAL ANALYSIS
There is a federal statute that defines when a drug or biologic is adulterated. It is 21 U.S.C. § 351. It has been on the books in some form since the Pure Food and Drug Act of 1906 and was codified into its current form in the 1938 Food, Drug, and Cosmetic Act. It is not obscure. Every first-year student of food and drug law learns this statute. It says, in plain English, that a drug is adulterated if it contains any substance that differs from its labeled or otherwise declared composition, or if it was prepared under conditions not in conformity with current good manufacturing practice. A product whose actual composition differs from what the manufacturer, the regulator, and the recipient were told it contained is, by statute, an adulterated product.
The COVID-19 mRNA products have been the subject of independent laboratory analysis since shortly after they entered the market. What those analyses show — and what Dr. Janci Lindsay’s team documented first and other teams subsequently replicated — is that the products contain SV40 promoter sequences and other residual DNA components that were not declared on the product label, not included in informed consent materials, and not acknowledged publicly by the regulator at the time of authorization. The regulator has since privately acknowledged the findings. The public has not been told.
CENSORING DR. JANCI’S TESTIMONY ON THE FOUND EVIL SV40 PLASMIDS!!!
A product whose actual composition differs from what the regulator approved and what the recipient was told — that is the statutory definition of adulterated. This is not a close call.
This is Part 5 of a series walking through Citizen Petition FDA-2025-P-1807. Parts 1 through 4 laid out the minimal risk waiver problem, the gene therapy classification problem, the DoD § 1520a predicate defect, and the Warp Speed authority problem. Part 5 addresses what is in the finished product — and what the statute says about a product with undisclosed components.
A. THE ADULTERATION STATUTE AND WHAT IT COVERS
21 U.S.C. § 351 defines adulteration across several provisions. The relevant texts for our purposes are —
“A drug or device shall be deemed to be adulterated — (a) Poisonous, insanitary, etc., ingredients … (1) If it consists in whole or in part of any filthy, putrid, or decomposed substance; or (2)(A) if it has been prepared, packed, or held under insanitary conditions whereby it may have been contaminated with filth, or whereby it may have been rendered injurious to health; or (B) if it is a drug and the methods used in, or the facilities or controls used for, its manufacture, processing, packing, or holding do not conform to or are not operated or administered in conformity with current good manufacturing practice to assure that such drug meets the requirements of this chapter as to safety and has the identity and strength, and meets the quality and purity characteristics, which it purports or is represented to possess.”
The statute has two operative hooks. One — if the product’s methods of manufacture do not conform to Current Good Manufacturing Practice. Two — if the product does not have the identity and strength, and meets the quality and purity characteristics, which it purports or is represented to possess. The second hook is the critical one. A product is adulterated when its actual composition diverges from the composition it claims, or claimed, to have. If the label says “mRNA encapsulated in lipid nanoparticles” and the finished product also contains residual DNA fragments including functional SV40 promoter sequences that were not disclosed, the statutory definition applies.
And these products are biological agents under federal statute, too
Adulteration is one frame. Biological agent is the other. Both apply to the same product.
Section 1520a defines a biological agent as “any micro-organism (including bacteria, viruses, fungi, rickettsiae, or protozoa), pathogen, or infectious substance, and any naturally occurring, bioengineered, or synthesized component of any such micro-organism, pathogen, or infectious substance.” The Biological Weapons Anti-Terrorism Act at 18 U.S.C. § 178 adds the harm criterion — any such agent “capable of causing death, disease, or other biological malfunction in a human, an animal, a plant, or another living organism.”
Read those definitions against the product. The COVID-19 mRNA shots are synthetic nucleic acid sequences encapsulated in lipid nanoparticles that instruct human cells to produce a spike glycoprotein. That is exactly what § 1520a means by “any bioengineered or synthesized component” of a pathogen. And the § 178 harm criterion is not theoretical. VAERS reports run orders of magnitude higher than for any prior vaccine. FDA’s own briefing documents acknowledged anticipated deaths before authorization. Spike-protein cardiovascular, neurological, and thromboembolic effects are in peer-reviewed literature. Sudden adult deaths, myocarditis in young athletes, pregnancy loss clusters, and persistent neurological injuries are in the public record.
Adulterated under 21 U.S.C. § 351. Biological agent under 50 U.S.C. § 1520a. Capable of causing death, disease, and other biological malfunction under 18 U.S.C. § 178. Three federal statutes. One product. One conclusion.
Why this overlap matters — when a product is both adulterated and a federally-defined biological agent capable of causing harm, the regulatory framework is supposed to escalate, not relax. FDA’s failure to enforce either framework here is what the petition asks the agency to correct.
B. WHAT DR. JANCI LINDSAY’S TEAM FOUND
Dr. Janci Lindsay is a toxicologist. She has extensive experience analyzing vaccine components. Beginning in 2023, her team conducted sequencing analysis on samples of commercially available COVID-19 mRNA vaccine vials. The analyses were performed using standard sequencing methodology. The findings were preserved, documented, and submitted to peer review.
Her team identified the presence of SV40 promoter sequences in the finished product. SV40 — Simian Virus 40 — promoter sequences are short segments of DNA that drive transcription of adjacent sequences at a high rate. They are used in molecular biology specifically when high expression is needed. Their presence in the final manufactured mRNA product is not theoretical. It is sequenceable. It is reproducible. It has been reproduced.
What makes SV40 promoter sequences regulatorily significant is that they are specifically used to facilitate integration of adjacent DNA sequences into the host genome. This is not a side effect — it is the engineered function of the promoter. When present in a biological product administered to a human, SV40 promoter sequences raise the clinical question of whether any residual DNA integrated near them has entered the recipient’s genome. That question has not been adequately answered because the monitoring framework — the 15-year gene therapy follow-up Part 2 laid out — was never implemented.
The SV40 promoter is not a contaminant in the sense of “accidental dust.” It is a functional genetic element with a known biological job — helping adjacent sequences get into the genome. Its undisclosed presence in an injected product is a regulatory question, not a rhetorical one.
C. INDEPENDENT REPLICATION
Dr. Lindsay’s findings have been replicated by independent laboratories in multiple jurisdictions. Dr. Kevin McKernan — a genomic scientist and former senior member of the Human Genome Project sequencing team — documented similar findings in his own published analyses. Health Canada has acknowledged the presence of residual DNA including SV40 promoter sequences in reviewed product samples, publishing partial regulatory findings that were subsequently limited in distribution. Regulatory bodies in Japan have received similar evidence from domestic toxicology teams.
What this means is that the findings are not a one-laboratory anomaly. They are consistent across independently sourced samples analyzed by independently funded teams using standard methodology. This is the threshold scientific evidence pattern a regulator is expected to take seriously. FDA has acknowledged the findings internally — per communications preserved in the petition record — while declining to acknowledge them publicly or to address them in regulatory guidance or in the agency’s response to the petition.
D. WHY SV40 PROMOTER PRESENCE MATTERS
To understand what the petition is asking FDA to address, you need to understand what SV40 promoter sequences actually do biologically. They are a tool used in molecular biology to drive transcription of adjacent DNA sequences at a high rate in mammalian cells. They are used because they work — they are robust, well-characterized, and produce reliable gene expression.
Their presence in the finished mRNA product creates two concurrent regulatory problems. First — their undisclosed presence means that neither regulators during review nor recipients at administration were informed that the product contained functional genetic elements designed to drive expression of adjacent sequences. Second — their biological function raises specific safety questions about genomic integration that the approved safety monitoring framework was not designed to surface. Dr. Lindsay’s research indicates that genomic integration can occur within six hours of exposure to the mRNA product. If that finding is correct — and it has been replicated — the implications for long-term safety, including oncogenesis risk and heritable genetic change, are the kind of implications that triggered the 15-year follow-up framework for gene therapy products in the first place.
E. THE INFORMED CONSENT BREACH
Even setting aside the safety questions, the informed consent framework requires disclosure of product composition to the recipient. Under the emergency use authorization statute at 21 U.S.C. § 360bbb-3(e)(1)(A)(ii), the Secretary is required to establish conditions ensuring that recipients are informed “of the significant known and potential benefits and risks of such use, and of the extent to which such benefits and risks are unknown.” The statute is explicit. Known composition elements are disclosed. Potential risks are disclosed. Unknown parameters are flagged as unknown.
The presence of SV40 promoter sequences was a known composition element at the manufacturer level — it had to be, because it was engineered into the production plasmid. It was not disclosed to recipients. Under the plain text of the statute, that is a failure of the informed consent framework. And under the Nuremberg Code and the binding international law obligations Part 6 will take up, failure to disclose composition to research subjects is itself an independent violation.
Recipients were not told the finished product contained functional genetic elements whose job in molecular biology is to drive expression of adjacent DNA. That is not a technicality. That is exactly the disclosure the informed consent statute was written to require.
F. DR. MICHAEL YEADON ON THE LIPID NANOPARTICLE AND SPIKE PROTEIN
Dr. Michael Yeadon’s expertise in pharmaceutical safety and development extends across decades at the highest levels of industry. He served as Vice President and Chief Science Officer for Pfizer’s Global Research and Development division. He co-founded Ziarco Pharma. His expert declarations attached to the petition address two distinct safety signals that run parallel to the SV40 promoter findings — the inherent toxicity of the spike protein produced by the mRNA product and the lack of adequate safety testing for the novel lipid nanoparticle delivery system.
Dr. Yeadon’s technical analysis is that the spike protein itself — as produced by transfected cells following mRNA administration — is biologically active in ways that include cellular signaling in cardiovascular and neural tissue, and that these signaling effects were not adequately characterized in the expedited pre-authorization review. His declarations form part of the scientific evidence base the petition asks FDA to engage on the record. FDA has not engaged them. Not in the response to the original 2022 petition. Not in the response to the January 2023 amendment. Not in any communication following the June 17, 2025 amendment.
G. VAERS AND WHAT THE DATA SHOWS
The Vaccine Adverse Event Reporting System is a passive surveillance system jointly maintained by FDA and CDC. VAERS data is a generation of adverse event reports filed by healthcare providers, manufacturers, and members of the public. It is not a causation-establishing system. It is a signal-generation system.
VAERS reports associated with COVID-19 vaccines are orders of magnitude higher than reports associated with any previous vaccine in the system’s history. This is not a political claim. This is the data in the database. The sheer volume of reports demands serious investigation under any reasonable reading of FDA’s post-market surveillance responsibilities. The agency’s response to this has been to cite VAERS’s acknowledged limitations as a reason to discount the signal. That response inverts the system’s purpose. VAERS exists to generate signals that are then independently investigated. Citing its limitations as a reason not to investigate is a disposition of the signal, not a process.
Internal FDA documents obtained through Freedom of Information Act requests — referenced in the petition and in the IOJ mandamus record — show that the agency was aware of multiple safety signals early in the vaccine rollout but chose not to communicate these risks to the public or to healthcare providers administering the vaccines. That FOIA-based finding is one of the factual foundations of the mandamus. The agency will be asked in federal court to explain the gap between what it knew and what it disclosed.
THE QUESTIONS FDA MUST ANSWER
1. Does FDA contest the presence of SV40 promoter sequences in commercially distributed COVID-19 mRNA vaccine vials? If so, on what evidentiary basis, given independent replication of Dr. Lindsay’s findings?
2. Were SV40 promoter sequences disclosed in the product labeling, the prescribing information, or the informed consent materials provided to recipients? If not, why not, in writing, on the docket?
3. Under 21 U.S.C. § 351, does a biological product containing functional genetic elements not disclosed to regulators during review meet the statutory definition of adulteration?
4. Has FDA independently analyzed samples of commercially distributed vaccine vials for residual DNA content, and if so, where in the administrative record are the sampling results?
5. Does the agency contest Dr. Lindsay’s finding that genomic integration can occur within six hours of exposure? If so, what counter-evidence does the agency rely on?
6. What, if any, post-market surveillance protocol does FDA intend to implement to address the integration risk the SV40 findings raise, given the absence of the 15-year gene therapy follow-up framework?
DID YOU KNOW
• SV40 — Simian Virus 40 — was first identified as a contaminant of early polio vaccines in the 1950s and 60s. Its historical presence in biological products is well-documented. Its use as an engineered promoter in molecular biology is also well-documented. The two uses are distinct but regulatorily related — both require disclosure.
• Dr. Kevin McKernan was on the original Human Genome Project sequencing team. His independent replication of Dr. Lindsay’s findings was published at anandamide.substack.com and has been cited in regulatory filings in multiple jurisdictions.
• Under 21 U.S.C. § 393(b), the Food and Drug Administration is required to protect public health by ensuring that drugs and biological products are “safe and effective.” Refusing to investigate replicated evidence of undisclosed genetic elements in a biologic is not consistent with the statutory mission.
• The statutory definition of adulteration at 21 U.S.C. § 351 does not require proof that the undisclosed component caused harm. It requires only that the product’s actual composition differs from its represented composition. That is the statutory test. It is already met.
THIS WEEK — YOU CAN HELP JUSTICE HAPPEN!
This is the fundraising week that carries this series to the federal courthouse. We need paid subscribers and monthly and one-time donations. That is the entire math — reader-supported, on a deadline, no corporate sponsors, no law firm behind us. If this work matters to you, this is the week to show up.
YOU CAN HELP IOJ FINALLY HOLD FDA ACCOUNTABLE. STOP THE SHOTS!
Share this series with a doctor, a lawyer, a reporter, a member of Congress, a state AG. The petition is public at regulations.gov/docket/FDA-2025-P-1807.
If you can pray on this one, please pray. If you can help donate it really helps. Sharing & comments help too! Whatever you can offer is the only thing that makes real justice happen in real time!
Part 6 about our Citizen Petition that Kennedy is lagging on is drafted. The International Law Dimension OF NO EXPERIMENTATION. ICCPR Article 7. Nuremberg Code. UNESCO Universal Declaration on Bioethics Article 6. The United States ratified these obligations and is bound by them. Part 6 walks through what the international legal framework actually requires and why the Charming Betsy canon forces domestic statutes to be read consistent with them. Learn how and why we will win!
When will these greedy genocidal criminals be arrested?
World law enforcement agencies must arrest Bill Gates, Walensky, Birx, Baric, Fauci, Dr. Peter Marks, Collins, Daszak and all former and present DOD, DARPA, CIA, CDC, NIH, WHO, FDA, AMA, AAP, doctors and hospital admin., HHS and big pharma and big tech. executives involved for crimes against humanity!
Fraud and Genocide are ...not included... in the total immunity from legal liability agreement under the PREP Act for the big Pharma criminals! I know, I've read it!
Nuremberg Code and RICO laws apply now! The Trump DOJ better wake the Fk Up and get busy!
Federal Crimes, Insurrection and conspiracy are what RICO laws are for!
Trial at 2 PM, Verdict ant 2:03, Executions at 2:27 PM. Rope is reusable!
You've been filing bullshit lawsuits for a decade;
Rosondich et al. v. Madison County et al., No. 1:15-cv-01120 (W.D. Tenn. May 13, 2015)
An expansive, sprawling civil rights complaint alleging a county-wide conspiracy and multiple constitutional violations. It did not survive initial judicial screening due to lack of factual support and legal barriers like immunity. Your allegations were "conclusory and not supported by specific, plausible facts." Court did you a favor by considering it because it "failed to meet basic pleading standards required in federal court."
The dismissal;
Rosondich et al. v. Madison County et al., No. 1:15-cv-01120, 2015 WL [unpublished] (W.D. Tenn. Aug. 20, 2015)



When will these greedy genocidal criminals be arrested?
World law enforcement agencies must arrest Bill Gates, Walensky, Birx, Baric, Fauci, Dr. Peter Marks, Collins, Daszak and all former and present DOD, DARPA, CIA, CDC, NIH, WHO, FDA, AMA, AAP, doctors and hospital admin., HHS and big pharma and big tech. executives involved for crimes against humanity!
Fraud and Genocide are ...not included... in the total immunity from legal liability agreement under the PREP Act for the big Pharma criminals! I know, I've read it!
Nuremberg Code and RICO laws apply now! The Trump DOJ better wake the Fk Up and get busy!
Federal Crimes, Insurrection and conspiracy are what RICO laws are for!
Trial at 2 PM, Verdict ant 2:03, Executions at 2:27 PM. Rope is reusable!
You've been filing bullshit lawsuits for a decade;
Rosondich et al. v. Madison County et al., No. 1:15-cv-01120 (W.D. Tenn. May 13, 2015)
An expansive, sprawling civil rights complaint alleging a county-wide conspiracy and multiple constitutional violations. It did not survive initial judicial screening due to lack of factual support and legal barriers like immunity. Your allegations were "conclusory and not supported by specific, plausible facts." Court did you a favor by considering it because it "failed to meet basic pleading standards required in federal court."
The dismissal;
Rosondich et al. v. Madison County et al., No. 1:15-cv-01120, 2015 WL [unpublished] (W.D. Tenn. Aug. 20, 2015)